Bringing a medical device or in vitro diagnostic device to the US market requires more than selecting a submission
route. Classification, intended use, predicate strategy, evidence, labelling, quality system readiness and post-market
obligations must support one another from the outset. A weakness in one area can delay review, create avoidable
questions or undermine the long-term viability of the US market strategy.
LFH Regulatory provides practical, coordinated FDA regulatory support for organisations at every stage of the
product lifecycle. Whether you are preparing your first US submission, expanding an established portfolio or
strengthening compliance after market entry, we can provide the specialist resource needed to define the pathway,
build the evidence, prepare the submission and respond effectively to the FDA.

Zara Malik
We assess the device, intended use, classification, product code, regulatory history and available predicates to identify
the most appropriate route. Where targeted FDA feedback would reduce uncertainty, we can help prepare for
Pre-Submission engagement and frame the questions that matter most to the development programme.
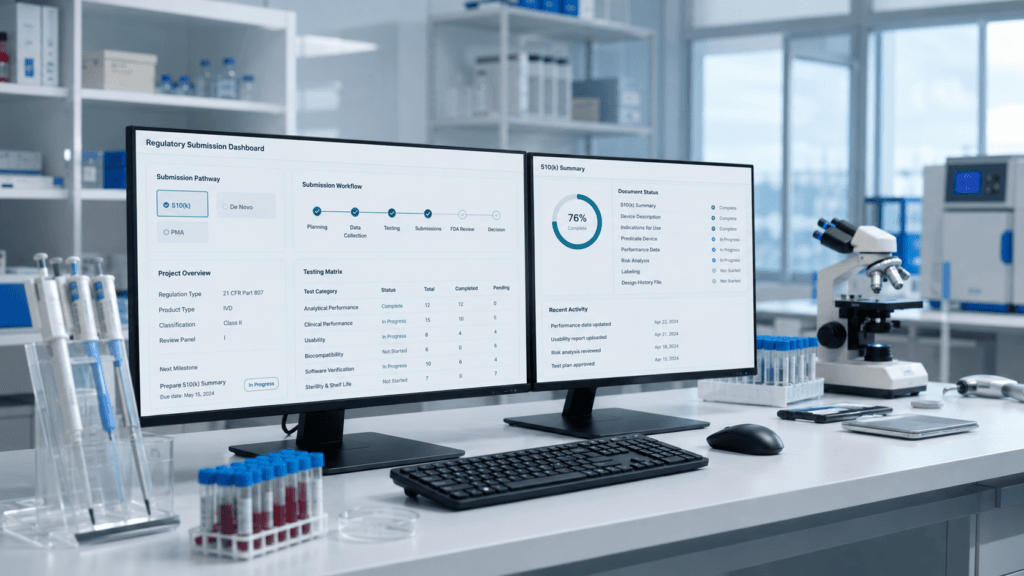
We translate the chosen pathway into a proportionate evidence and documentation plan. This may include risk
management, bench testing, biocompatibility, electrical safety, software and cybersecurity, sterilisation, shelf life,
usability, clinical evidence and labelling, depending on the device and the questions the FDA must resolve.
US compliance continues after clearance, classification or approval. We help organisations prepare for ongoing
quality, registration, listing, reporting and post-market obligations, and assess whether product, process or labelling changes create new regulatory requirements.
A strong US strategy starts with a defensible understanding of the product and its regulatory status. We can support
device classification, product code research, predicate and precedent analysis, intended use review, pathway selection, evidence planning, submission sequencing and the identification of risks that could affect development or market
entry.
When key questions cannot be resolved confidently through existing regulations, guidance and precedent, early FDA
engagement can provide valuable direction. We can help define the objectives of a Pre-Submission, prepare the briefing package, develop clear questions, support meeting preparation and translate the FDA’s feedback into
practical actions for the development and submission teams.
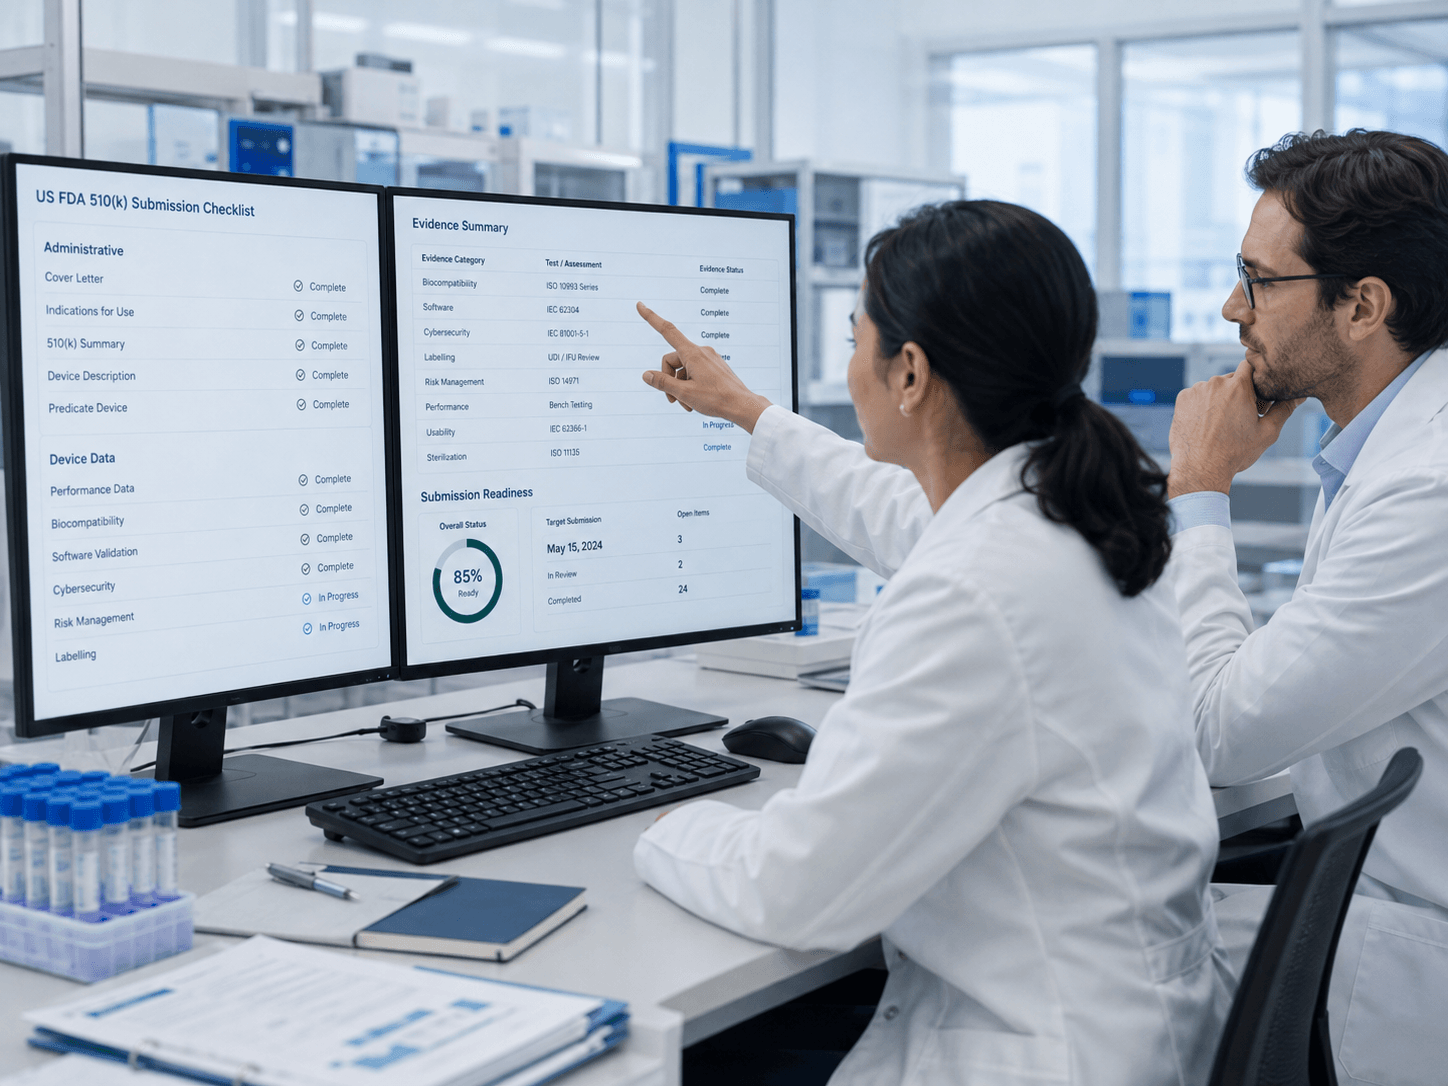
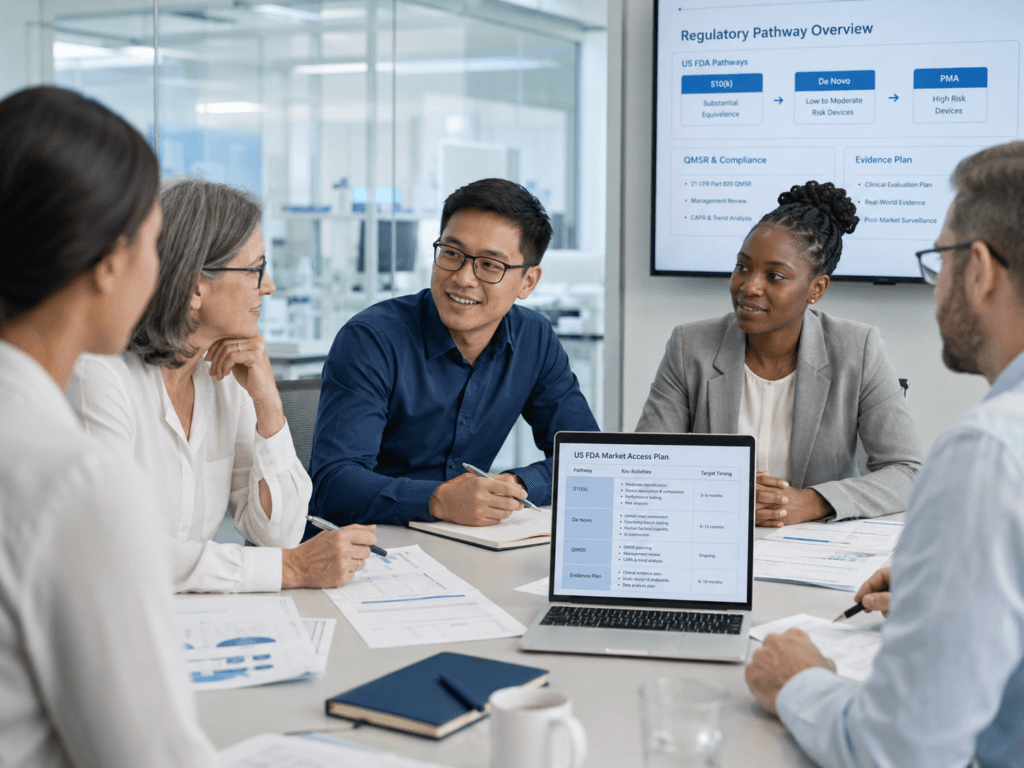
For devices requiring a 510(k), the submission must demonstrate substantial equivalence to an appropriate legally marketed predicate. We can support predicate selection, substantial equivalence strategy, gap assessment, testing plans, documentation development, eSTAR preparation, quality review, submission management and responses to FDA requests for additional information. Our aim is to create a coherent submission in which the intended use, technological comparison, performance evidence and labelling tell the same regulatory story.
The appropriate US route depends on the device classification, intended use, technological characteristics, available predicates and level of risk. A product may be exempt from premarket notification, require a 510(k), qualify for the De
Novo pathway or need Premarket Approval. The correct decision must be supported by a documented classification and evidence strategy, not an assumption based on a superficially similar product.
For manufacturers based outside the United States, market access also involves requirements beyond the premarket
submission. Quality Management System Regulation compliance, establishment registration, device listing, US Agent
arrangements, labelling, complaint handling, reporting and post-market controls must be considered as part of one
connected plan. Early coordination helps prevent duplication, identify evidence gaps and keep commercial
expectations aligned with regulatory reality.
The De Novo pathway may be appropriate for a novel device without a suitable legally marketed predicate when
general controls, or general and special controls, can provide reasonable assurance of safety and effectiveness. We can help assess pathway suitability, define the proposed classification and special controls, plan the evidence package, prepare the request and support interactions with the FDA throughout review.
Premarket Approval is the FDA’s most stringent device marketing application and is generally required for Class III
devices. A PMA must contain valid scientific evidence that provides reasonable assurance of safety and effectiveness for the intended use. We can support regulatory planning, evidence strategy, clinical and non-clinical documentation, application structure, quality review, submission coordination and responses during the FDA review process.
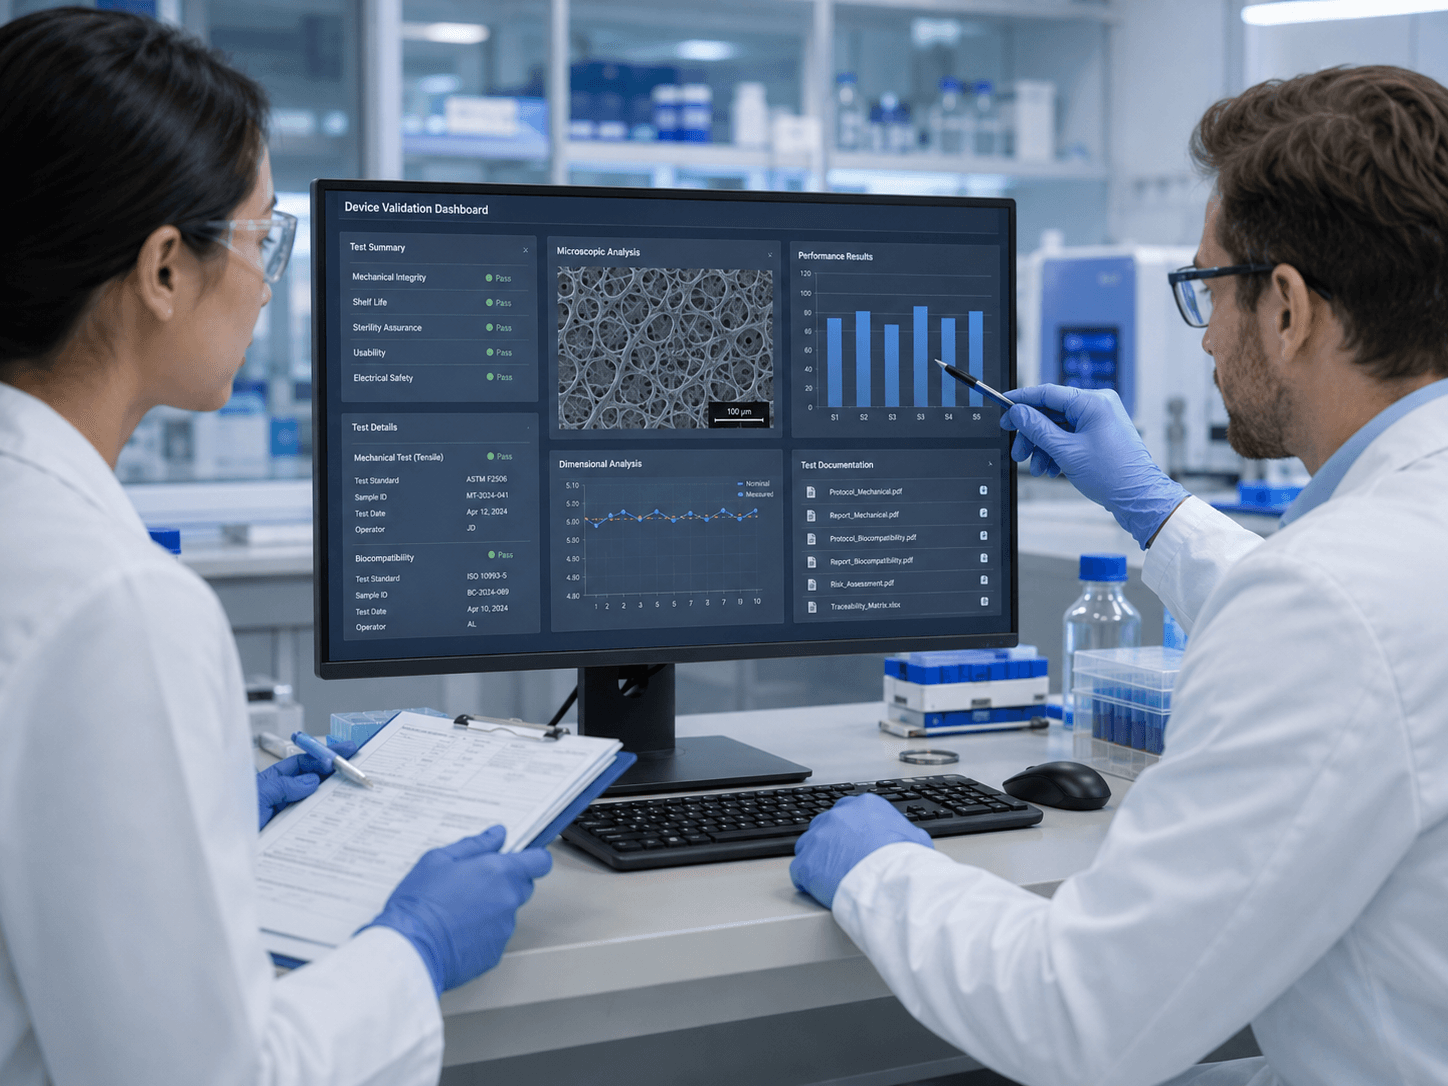
The evidence required is specific to the device, claims, risk profile and regulatory pathway. We can help identify
applicable standards and guidance, assess existing data, define gaps and coordinate the strategy for bench, biocompatibility, electrical safety, software, cybersecurity, sterilisation, packaging, shelf-life, human factors and
clinical evidence. Where specialist studies are required, the work can be coordinated with appropriate technical and clinical partners.
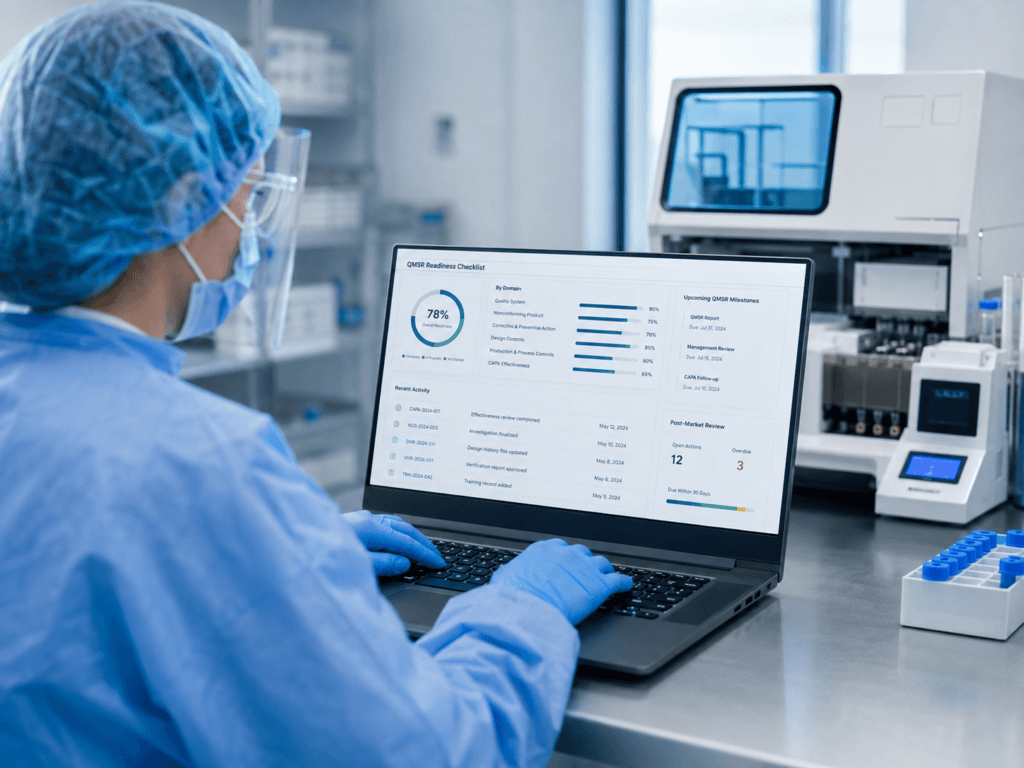
The FDA’s Quality Management System Regulation became effective on 2 February 2026 and incorporates ISO
13485:2016 by reference, while retaining FDA-specific legal and regulatory requirements. We can support QMSR
readiness assessments, gap analysis, procedure development, design and development controls, risk management integration, audit preparation, remediation planning and inspection readiness. This allows organisations already operating an ISO 13485 quality management system to build on existing controls while addressing the requirements that remain specific to the United States.
Manufacturers must understand which establishments require registration, which devices must be listed and when a foreign establishment needs a designated US Agent. We can support the preparation and coordination of registration and listing activities, review the information for consistency with the authorised device and help maintain the records as products, establishments or responsibilities change.
A technically sound submission must also be complete, navigable and presented in the format required by the FDA. We can support document planning, compilation, formatting, hyperlinking, administrative content, reviewer guides, quality control and electronic submission preparation, including eSTAR where applicable. Clear ownership and version control help reduce last-minute errors and keep internal reviewers working from the same approved information.
Add experienced regulatory and quality capability to your team without committing to a fixed internal structure. We
can take responsibility for a defined work package or provide ongoing support as priorities evolve.
We work closely with internal stakeholders, technical specialists and external partners, keeping responsibilities, decisions and next actions clear. Our approach is structured, proportionate and focused on helping the programme move forward.
Regulatory strategy, evidence, quality systems, submissions and post-market compliance are connected. Maintaining
continuity across these areas reduces handover risk and helps organisations make better-informed decisions as the
product and regulatory landscape develop.
Speak with LFH Regulatory to discuss your device, intended use, current evidence and the specialist support needed to move forward with confidence.